Yes, relying on a single cryocooled protein structure can mislead parts of computational structure-based drug design. Cooling crystals for X-ray data collection can shift the conformations captured in the structure, including side chains, ligand poses, solvent networks and allosteric states. That matters when a model is treated as the definitive shape of a binding site or used to calibrate and validate computational predictions. It does not mean cryogenic structures are inherently unreliable: they remain valuable, but their suitability depends on the biological question and the modeling task.
Why a temperature-conditioned structure can affect drug design
A crystal structure is a measurement, not the protein’s complete ensemble
Proteins occupy multiple conformations. An X-ray structure represents states visible under particular experimental conditions, rather than every state the protein samples in solution or in a cell. Cryocooling helps limit radiation damage during data collection, but it can also shift conformational populations. A model derived from a cryogenic structure may therefore omit or underrepresent a state that matters for ligand binding or protein function.
As an Amazon Associate I earn from qualifying purchases.
Where the difference can enter a computational workflow
- Docking and pose interpretation: a pocket’s shape, side-chain rotamers, or nearby solvent can differ with temperature, changing which poses appear plausible.
- Virtual screening: a candidate may fit the conformation captured in the structure without fitting other relevant conformations, or a useful binding state may be absent from the model.
- Calibration and validation: methods assessed against structures that encode temperature-dependent features may be judged against a reference that does not represent the state relevant to the intended application.
- Allostery: a structure that does not reveal a conformational network or alternate state may provide an incomplete basis for interpreting how a ligand affects a distant site.
These are risks demonstrated in particular structural comparisons, not evidence that a specific docking score is systematically wrong or that cryogenic structures cause a known rate of failed drug-discovery predictions. Bradford and colleagues’ study describes temperature artifacts as a concern for computational calibration, validation and ligand discovery.
What’s actually slowing this PC down?
Pick the symptom - the matching free tool is one click away.
What comparisons between cryogenic and room-temperature structures found
A 30-protein comparison found widespread side-chain changes
In a 2011 analysis of 30 proteins, Fraser and colleagues found that crystal cryocooling remodeled the conformational distributions of more than 35% of side chains in the structures they examined. In H-Ras, room-temperature electron-density maps showed an allosteric network that was not apparent in cryogenic maps; the network was consistent with solution NMR observations. The result demonstrates that cooling can alter what is visible in structural data. The reported percentage describes that study’s protein set, not a universal estimate for any new structure. Read the Fraser et al. study.
#1 Best Overall

- FOR BASIC TEACHING TO ADVANCED SCIENCE: 444 pieces molecular model kit, including 136 atoms, 158 bonds and 150 parts for Carbon-60(Fullerene), provides to students from Grade 7 to Graduate level.
- TWO CHEMICAL STRUCTURE MODELS: The ball-and-stick models use spheres to represent atoms and sticks to represent chemical bonds. In the space-filling model, the spheres are drawn to scale and are next to one another as atoms are in real molecules.
- CHEMISTRY EDUCATIONAL MOLECULE MODEL IN 3D: It can display chemical structure, molecular bond, and bond angle in all directions. Demonstrate fundamental molecular geometry, chemical molecular structure, stereochemistry with 3D modeling studies.
- EASY TO LEARN: The universal standard adopted for each atom's color makes it easier for you to use and learn. Atoms and chemical bonds combine tightly and firmly and can be easily disassembled by disconnecting tools.
- If you’re not in love with it for whatever reason, we’ll give you a full replacement or refund—no questions asked. If you have any doubt, please tell us. With nothing to worry about, or even to share with your friends, try it now.
T4 lysozyme showed a ligand-relevant apo conformation
Bradford and colleagues studied T4 lysozyme L99A, a well-characterized cavity system, along with additional protein classes. Their comparison found a room-temperature apo helix conformation relevant to ligand binding that was hidden in the cryogenic structure, as well as temperature-dependent differences in side chains and ligand poses. This shows how a cryogenic model can leave out a binding-relevant conformation in a tested system; it does not establish how often that happens across targets.
PTP1B fragment screens produced different structural pictures
A 2023 PTP1B study compared two room-temperature crystallographic fragment screens with an earlier cryogenic screen that included many of the same fragments. The room-temperature work reported fewer and often weaker binding observations, but also revealed unique poses, altered solvation, new binding sites and distinct allosteric responses. The comparison shows that collection temperature can affect both apparent fragment hits and the structural interpretation of binding. It is specific to PTP1B and the study’s experimental design, not a general prediction that room-temperature screens will produce fewer hits. See the PTP1B study.
Rank #2
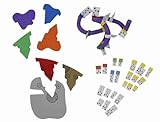
- Two popular kits combined in a version perfect for one student in a tutoring center or home study
- Identify and sort the side chains based on their chemical properties
- Explore primary, secondary, and tertiary protein structure
- Fold a zinc finger protein motif with alpha helices and beta sheets after calculating the scale
- Model active sites, mutations, denaturation, and reverse engineering
What cryogenic and room-temperature crystallography each offer
Neither temperature is best for every target. Cryogenic data collection can make it more practical to limit X-ray damage and obtain complete, high-resolution datasets. Room-temperature crystallography can preserve or expose conformational states that cooling shifts, but crystal damage and sample throughput can make data collection more demanding.
| Consideration | Cryogenic crystallography | Room-temperature crystallography |
|---|---|---|
| Radiation damage and data collection | Cooling limits X-ray damage and helps enable complete, high-resolution datasets, as described in Chemistry World’s 2021 report. | For many proteins, crystals can deteriorate rapidly during data collection. Keith Wilson, a crystallography methods expert at the University of York, told Chemistry World that collecting a complete dataset can require many crystals. |
| Conformational information | Cooling may shift conformational populations or make some states less visible, as observed in the comparative studies above. | Can reveal conformational states and networks not apparent in cryogenic structures, though results depend on the target and experiment. |
| Best use in modeling | Useful structural evidence, but a single cryogenic model should not automatically be assumed to represent every state relevant to binding or function. | A complementary view when the question depends on flexible regions, transient pockets, ligand poses, solvent or allostery. |
The methods literature treats room-temperature X-ray crystallography as an approach to optimize, not a universal replacement for cryogenic data collection. The 2023 IUCrJ review discusses methods and future optimization.
Rank #3

- Explore the structure of proteins and function of enzymes
- Construct the double helix structure of DNA
- Examine nucleotide sequences and the replication, transcription, and translation of DNA
- Connect the molecular to the microscopic structures of genetic information
- Examine the phospholipids, cell membranes, and transport proteins
How to use cryocooled structures more carefully
- Match the structure to the question. Decide whether the task is to explain a known ligand pose, search for candidate binders, model a flexible pocket, or study allosteric effects. A structure useful for one purpose may not capture the states needed for another.
- Check experimental conditions before interpreting fine details. Record whether the structure was collected cryogenically or near room temperature, and pay particular attention to flexible loops, side chains bordering the pocket, ligand orientation and nearby solvent.
- Look for relevant alternate states. If a binding site or mechanism depends on conformational flexibility, compare available structures or use ensemble-sensitive evidence rather than treating one coordinate model as the only possible protein shape.
- Validate against the intended use. Where feasible, test whether conclusions persist across relevant structural states or are supported by independent experimental evidence. A prediction that depends on one temperature-conditioned feature deserves more caution than one that holds across multiple lines of evidence.
- Keep the limits explicit. When reporting a computational result, identify the structural model and its collection context. Do not present a pose or score as proof of binding or mechanism solely because it fits one crystal structure.
What the evidence does not show
The comparative studies establish that cooling can change structural observations and that those changes can matter for ligand interpretation and computational workflows. They do not show that every cryogenic structure is misleading, that room-temperature structures are always more biologically representative, or that replacing cryogenic structures would improve every prediction. Nor do they quantify a universal loss of docking accuracy, prospective hit rate or clinical success. The practical conclusion is narrower: treat a cryocooled structure as valuable evidence whose representativeness should be assessed for the specific modeling question.
Quick Recap
Best Value

- (C) FromSoftware, Inc. All rights reserved.
- Introducing the Armored Core series from the Square Enix Plastic Model Assembly Kit Series, Structure Arts (Structure Arts) series
- From ARMORED CORE comes the lineup of Chrome XCA-00 Unfang
- From ARMORED CORE2, the Geo Matrix ZCL-XA/2, Emroad ECM-XR00 lineup
- Based on the supervision of the From Software Company, each unit is reproduced in high detail
Rank #4

- Easy to learn and handle: The molecular model kit is simple to assemble, store, and carry. Atoms connect securely yet can be easily detached using the included disconnect tool, making it perfect for repeated classroom use.
- Practical for teaching: This model helps students visualize the structural features of protein molecules covered in textbooks. It boosts learning interest and allows teachers to explain complex concepts more clearly and intuitively.
- Versatile building options: The protein molecular model can be used to build both common and slightly complex protein structures, making it suitable for teaching demonstrations and laboratory experiments.
- 3D visualization aid: With this 3D modeling kit, students can explore molecular structures and angles from every angle, gaining a deeper understanding of molecular geometry and spatial relationships.
- Bright and color-coded: The model includes colorful atoms and connectors that follow standard color conventions, making identification easier. The vibrant colors and quality construction keep students engaged and simplify learning.
Product prices and availability are accurate as of the date/time indicated and are subject to change. Any price and availability information displayed on Amazon at the time of purchase will apply.




